
Updated:
Informed consent forms are signed agreements to join a research study. For federally supported studies, 45 CFR 46.116(b) lists what the form explains: purpose, risks, benefits, withdrawal. Researchers and students can fill out the free template below or print the PDF.
Build Your Document
Answer a few simple questions to make your document in minutes
Save and Print
Save progress and finish on any device, download and print anytime
Sign and Use
Your valid, lawyer-approved document is ready
An informed consent form for research is the document a volunteer reads and signs before joining a study, after the researcher explains the study's purpose, what the volunteer will do, and the risks and benefits of taking part.
The form is the written end of a longer conversation. The researcher talks the volunteer through the study, answers questions and gives them time to decide. The signed form then shows that the volunteer agreed, and on what terms.
A completed form holds:
Research teams also call it an ICF, a consent document or a research consent form. Hospitals, universities, survey teams and companies that run studies all use one, and many give their researchers a house template to start from. For consent outside research, such as a school trip or a photo release, see the other consent document templates.
The federal rules in 45 CFR 46.102 call the people who take part human subjects; most consent forms simply say participants.
This short glossary explains the words you will see most often on a research consent form.
You need an informed consent form when a study enrolls people and its rules call for signed consent; under 45 CFR 46.117(a), a Common Rule study documents consent with a written form approved by the IRB and signed by the participant or a legally authorized representative.
The Common Rule covers research involving human subjects that is conducted, supported or otherwise subject to regulation by a federal department or agency that has made the policy applicable to it, as 45 CFR 46.101(a) sets out.
Clinical investigations regulated by the Food and Drug Administration follow their own consent rules in part 50, whose scope is set by 21 CFR 50.1.
Universities, hospitals and journals can set their own consent policies too. Ask your IRB or ethics office which ones apply to your study before you write the form.
An IRB may waive the signed consent form for some or all participants, but only if it finds any of the conditions listed in 45 CFR 46.117(c).
Researchers running online surveys or phone interviews often ask the IRB about this option. Until the IRB says yes, plan on a signed form.
Some studies are exempt from the Common Rule: under 45 CFR 46.104(a), research whose only involvement of human subjects falls in one or more of the categories in 46.104(d) is exempt, except for the conditions each category sets.
One category, 45 CFR 46.104(d)(2), covers research that only includes educational tests, survey procedures, interview procedures or observation of public behavior, and only if at least one of its listed criteria is met, for example that the information is recorded so the participants cannot readily be identified.
Ask your IRB or the institution's human research office whether your study qualifies before you drop the signed form. An exempt study may still give participants a short information sheet about the study.
Virginia is one state with a human research statute of its own, Va. Code § 32.1-162.18, and the rules elsewhere differ.
Under Va. Code § 32.1-162.18, informed consent for human research in Virginia is subscribed in writing and witnessed when the subject can make an informed decision; a legally authorized representative signs, with a witness, when the subject cannot; and both the minor and the representative sign for a minor otherwise capable of consenting.
Va. Code § 32.1-162.18 also bars any consent form language through which the subject waives or appears to waive legal rights, including a release from liability for negligence.
These Virginia rules do not reach every study: Va. Code § 32.1-162.20 exempts human research that is subject to federal human subjects policies and regulations from that chapter.
Ask your IRB which rules apply where the study runs, beyond the federal ones on this page.
An informed consent form for a federally supported study covers the basic elements listed in 45 CFR 46.116(b), from the purpose and procedures of the study to its risks, its benefits and the participant's right to stop at any time.
The table starts with the key information rule of 45 CFR 46.116(a)(5)(i), then follows the order of 45 CFR 46.116(b) and says, in plain words, what to write for each element.
| Part of the form | What to write in it | Source |
|---|---|---|
| Key information | Begin the form with a concise and focused presentation of the key information that helps a person see why they might or might not want to take part. | 45 CFR 46.116(a)(5)(i) |
| Research, purpose, length and procedures | Say that the study is research, why it is done, how long the participant takes part and what they will do; point out any procedure that is experimental. | 45 CFR 46.116(b) |
| Risks and discomforts | Describe the reasonably foreseeable risks or discomforts. | 45 CFR 46.116(b) |
| Benefits | Describe the benefits to the participant or to others that can reasonably be expected. | 45 CFR 46.116(b) |
| Alternatives | Name other procedures or treatments that might help the participant, if any exist. | 45 CFR 46.116(b) |
| Confidentiality | Explain how far records that identify the participant will be kept confidential. | 45 CFR 46.116(b) |
| Compensation and injury | Only for research with more than minimal risk: say whether there is compensation and whether medical treatment is available if an injury occurs. | 45 CFR 46.116(b) |
| Contacts | Say whom to contact with questions about the research and about participants' rights, and whom to contact after a research-related injury. | 45 CFR 46.116(b) |
| Voluntary participation | Say that taking part is voluntary, that saying no brings no penalty or loss of benefits, and that the participant can stop at any time without penalty. | 45 CFR 46.116(b) |
| Identifiable information and specimens | Only when the study collects identifiable private information or identifiable biospecimens: say whether, with identifiers removed, they could be used in future research. | 45 CFR 46.116(b) |
FDA-regulated trials use a near-identical list in 21 CFR 50.25(a); its confidentiality element also notes that the Food and Drug Administration may inspect the records.
Some studies add more items, such as unforeseeable risks, costs to the participant, or what happens when someone leaves early. Your IRB's own template shows which extra items your study takes.
Write every part for the reader, not for the review board. Short sentences, everyday words and a clear heading for each part make the form easier to read, and easier to ask questions about.
A clinical trial consent form follows the Food and Drug Administration's consent rules when the trial is a clinical investigation the agency regulates, which 21 CFR 50.1 brings under part 50.
Under 21 CFR 50.27(a), the participant or a legally authorized representative signs and dates the form at the time of consent, on a written form approved by the IRB.
The person who signs gets a copy, under the same section, 21 CFR 50.27(a).
For a trial with more than minimal risk, the form explains whether compensation is offered and whether medical treatment is available if an injury occurs, as 21 CFR 50.25(a) sets out.
For an applicable clinical trial as defined in 42 U.S.C. 282(j)(1)(A), 21 CFR 50.25(c) requires the consent documents to give a set statement that a description of the trial will be available on ClinicalTrials.gov. Copy that statement word for word from 21 CFR 50.25(c); do not reword it.
Trial forms tend to run long. When the trial is also covered by the Common Rule, the form begins with a concise and focused presentation of the key information, under 45 CFR 46.116(a)(5)(i). Plain words and a clear list of visits and tests also help a volunteer see what the trial asks of them.
The FormsPal template fits smaller studies best. A sponsor or hospital running a drug or device trial usually hands each site a consent template of its own, and the trial team fills that in.
Yes, a participant can withdraw at any time: under 45 CFR 46.116(b), the form says that taking part is voluntary and that the participant may stop at any time without penalty or loss of benefits they are otherwise entitled to.
Consent also does not end at the signature. The participant can ask the research team questions during the study and after it ends.
A good form tells the participant how to withdraw: whom to tell, whether a phone call or an email is enough, and what happens to data or samples already collected.
Many forms also say when the researcher may end a person's part in the study, for example when they no longer meet the study criteria or cannot follow the study procedures.
Follow the steps below to complete the informed consent form for a research study. Open the template, then work from the top of the document down to the signature lines, one part at a time.
Have the study plan and your IRB's approval details at hand before you begin.
The screenshots under each step show the FormsPal Informed Consent template filled with sample entries in blue, so you can see where each answer goes. The names, addresses, dates and amounts are examples only.
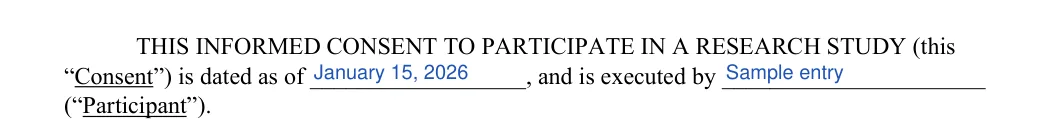
Date and Participant
Find the opening paragraph on page 1. Enter the date of the consent and the name of the participant.
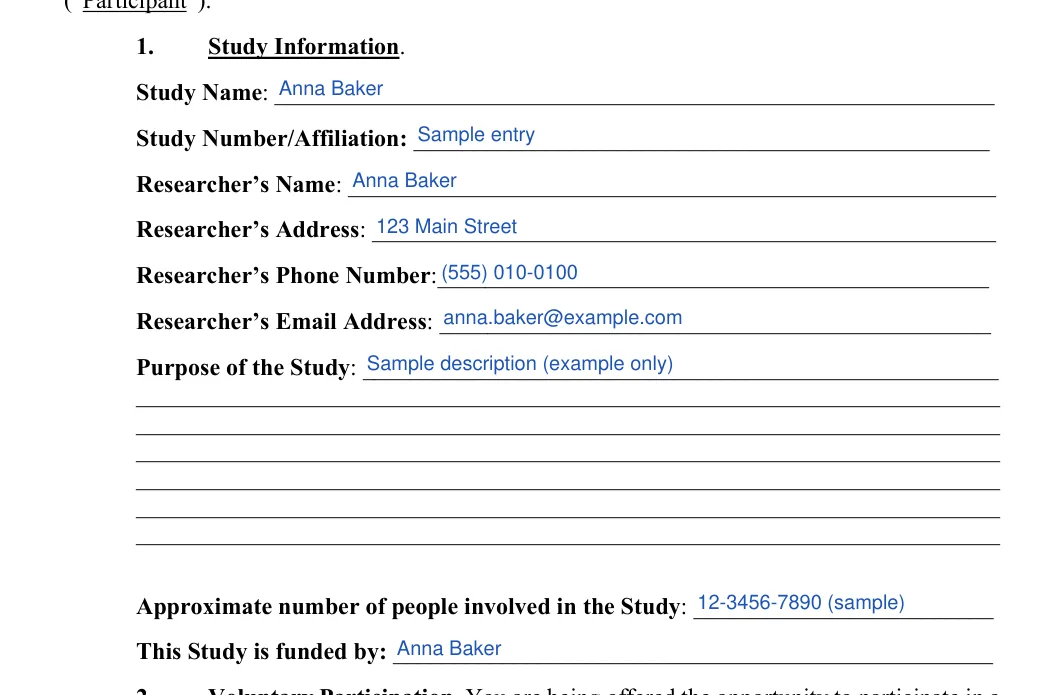
Study Information
Next, describe who runs the study and how to reach them.
Fill in the study name, the study number or affiliation, and the researcher’s name, address, phone and email. Add a short purpose, the approximate number of people involved and who funds the study.
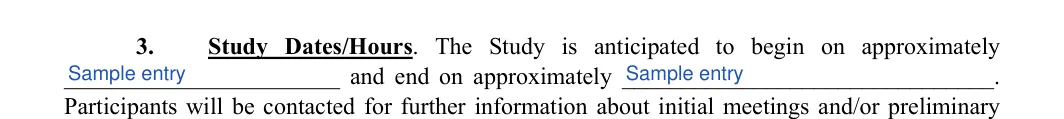
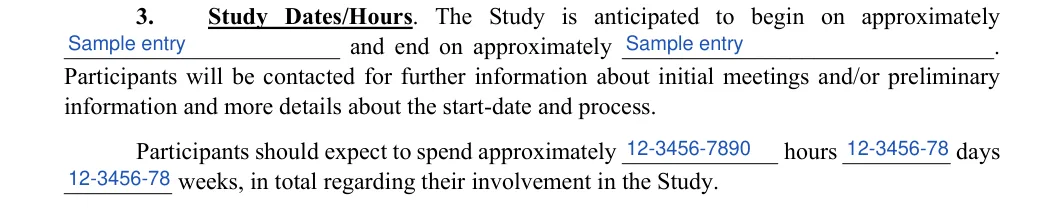
Study Dates
Give the period when the research runs, so the participant knows when it starts and ends.
At the top of page 2, write the approximate begin and end dates.
Study Hours
Tell the participant how much of their time the study takes in total.
Still in the Study Dates and Hours part, enter the total hours, days and weeks.
Participant Criteria and Purpose of the Study
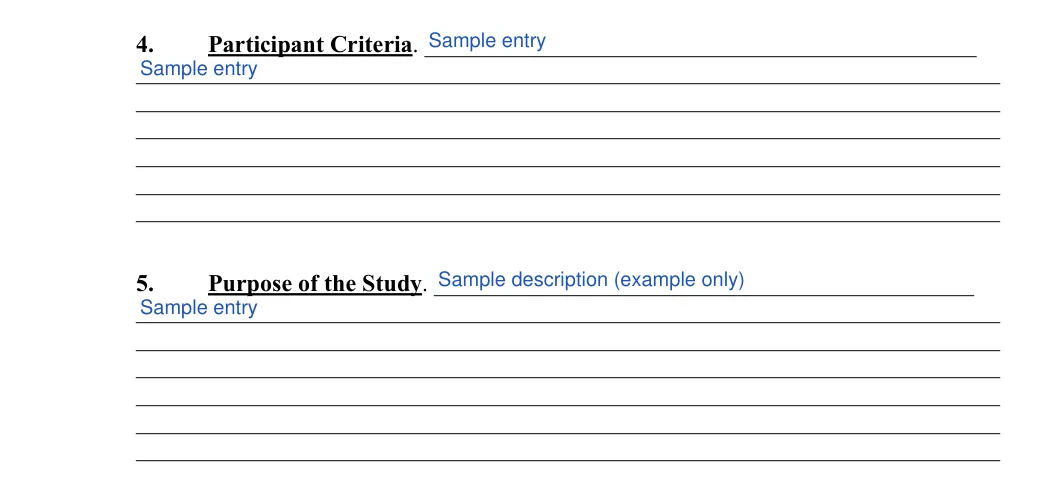
On page 2, find the Participant Criteria heading. Describe who can join the study and who cannot, in plain words.
Then write a short purpose under the Purpose of the Study heading.
Risks and Benefits of Involvement
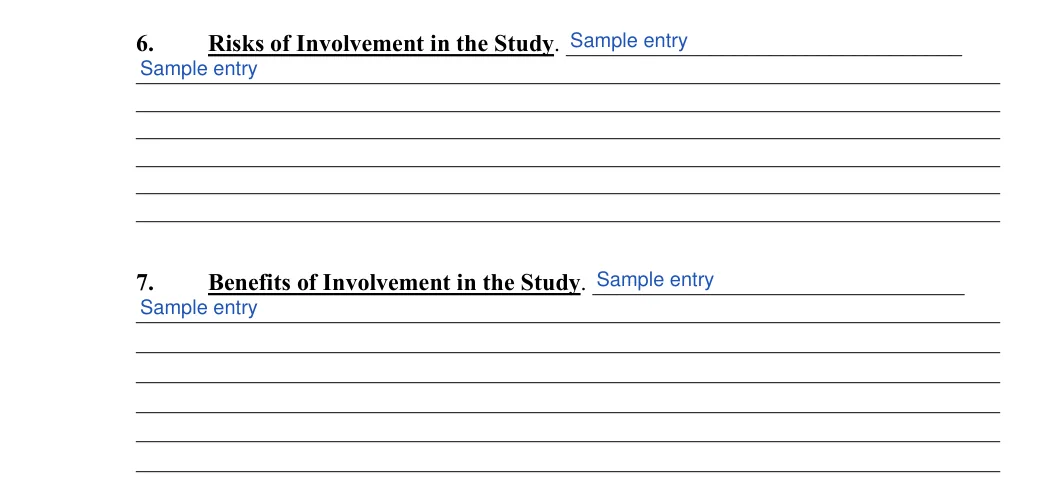
List each risk or discomfort you can reasonably foresee, small ones such as tiredness included, and describe the benefits without overstating them.
Write the risks first and the benefits next, still on page 2.
Compensation, Costs and Confidentiality
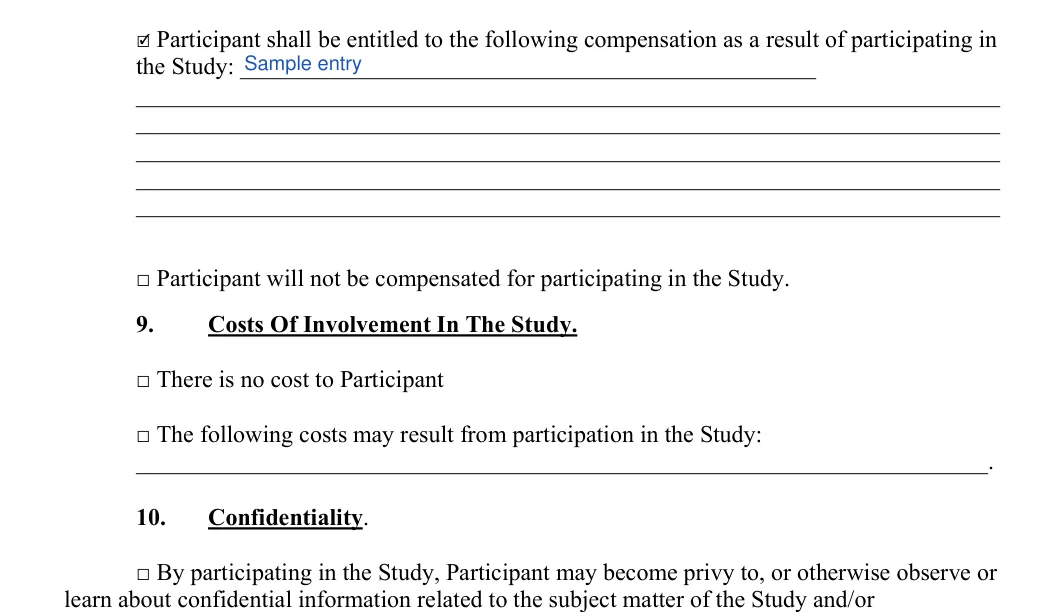
Say whether participants are paid for taking part and whether they pay for anything themselves, such as travel or treatment.
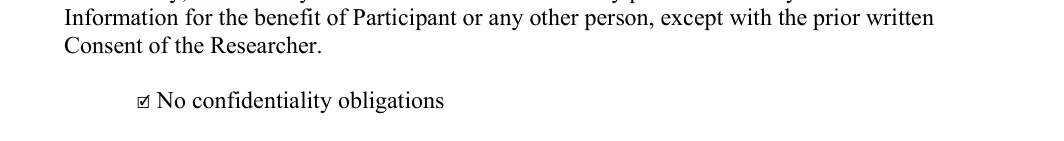
On page 3, tick the compensation option and write in the amount if one applies. Tick whether there is no cost or list the costs. Then tick the confidentiality paragraph if it applies to the study.
No Confidentiality Obligations
Every form says how far records that identify the participant are kept confidential, a basic element under 45 CFR 46.116(b). Tick the No confidentiality obligations box only when that is the true answer for the study, for example when it keeps no records that identify anyone, and add a plain line saying so.
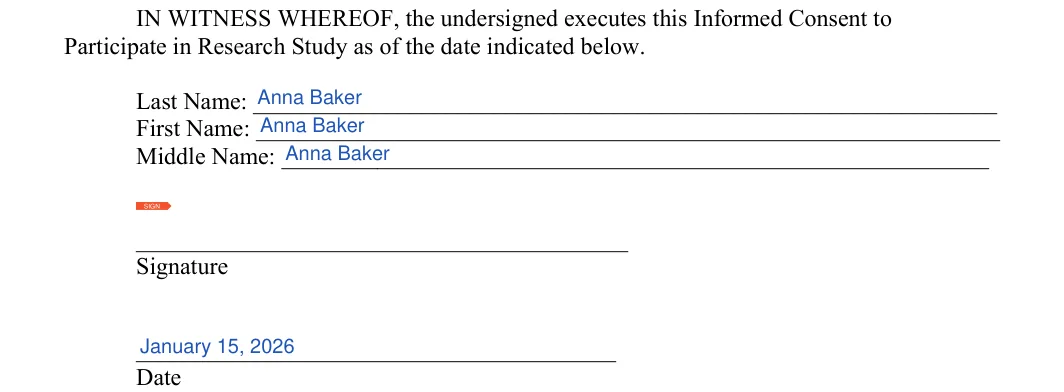
Name, Signature and Date
The participant signs only after reading the whole form and getting answers to their questions.
Under the closing line on page 3, print the last, first and middle name, sign on the Signature line and write the date.
Give the signer a copy afterwards; 45 CFR 46.117(a) and 21 CFR 50.27(a) both call for one.
The nine parts above do not hold every element of the table in the section on what to include. Under 45 CFR 46.116(a)(5)(i) and 45 CFR 46.116(b), add these in your own words before IRB review, where they apply to your study:
The informed consent form example below shows how the parts of the template read once filled in for a small, made-up survey study, so you can see the tone and the level of detail that fit a real form.
Sample only: a made-up study, not a real form.
Study title: Sleep Habits of College Students
Researcher: Dr. Jane Sample, Department of Psychology, Example University, jane.sample@example.edu
Purpose: This research study looks at how students sleep during exam weeks.
What you will do: You will answer an online questionnaire of 30 questions. It takes about 20 minutes.
Risks: Some questions about sleep and stress may feel personal. You can skip any question.
Benefits: There is no direct benefit to you. The results may help the university plan student health services.
Payment: You will receive a $10 gift card after you finish.
Privacy: Your answers are stored without your name on a password-protected computer.
Your choice: Taking part is voluntary. You can stop at any time without penalty.
Questions: About the study, contact Dr. Jane Sample. About your rights as a participant, contact the Example University IRB.
Agreement: I have read this form, my questions were answered, and I agree to take part. Name ______ Signature ______ Date ______
Replace every line with your own study's details. Never carry the sample's risks, payment or contacts into a real form.
The same consent form works for survey, psychology, quantitative and case study research; what changes is the detail in the procedures, risks and confidentiality parts, which should match what each kind of study asks of the people in it.
An informed consent form for survey research tells people how long the survey takes, whether they can skip questions, and whether their answers are anonymous or linked to a name or email address.
For an online survey, the consent text often sits on the first screen, with an agree button in place of a signature. That setup needs either an IRB waiver of the signed form under 45 CFR 46.117(c) or a finding that the study is exempt under 45 CFR 46.104, as an anonymous survey may be under 45 CFR 46.104(d)(2). Your IRB or ethics office tells you which applies.
An APA-style informed consent form is a psychology consent form laid out in the format a psychology course or department asks for; its content follows the same parts as the template on this page.
Ask your instructor or IRB for the department's own template before you start. If there is none, start from the FormsPal form and add the statements it does not hold, listed under the filling steps. Describe each task the participant will do, in the order they will do it.
A consent form for quantitative research explains what data is collected, how it is stored and who can see it, and whether results are reported only as group figures that name no one.
A case study consent form asks permission to describe a single person, family or organization in detail. Say what will be published, whether names or photos appear, and whether the participant can review quotes before release.
These short answers cover questions people often ask about an informed consent form for a research study. If you still have a question about your own study, ask the researcher or the institution that runs it.
Yes, a legally authorized representative can sign for a participant who cannot consent: 45 CFR 46.117(a) lets either the subject or the subject's legally authorized representative sign.
Under 45 CFR 46.102, that is a person or body authorized under applicable law to consent on the participant's behalf, so who qualifies depends on the law that applies where the study runs.
When children take part, the researcher seeks a parent's or guardian's permission under 45 CFR 46.408(b), and the child's own assent when the IRB judges the child capable of giving it, under 45 CFR 46.408(a).
For studies under 45 CFR 46.406 or 46.407, 45 CFR 46.408(b) calls for both parents' permission unless one parent is deceased, unknown, incompetent or not reasonably available, or only one parent has legal responsibility for the child's care and custody.
For studies under 45 CFR 46.404 or 46.405, the IRB may find one parent's permission sufficient, 45 CFR 46.408(b).
IRB stands for institutional review board. It is a group at a hospital, university or research center that reviews the plan for a study. If your form names an IRB, you can contact it with questions about the study.
Yes, for a Common Rule study: 45 CFR 46.117(a) counts a signature made in an electronic format. Check with the IRB which e-signature tools it accepts.
Yes. You have two ways to do it:
Ask the researcher or the study team about anything on the form that you do not understand. Do this before you sign.
General information, not legal or tax advice.
